
On the way to the EU medical device market there is a role that often works in the background but is indispensable: the European Authorised Representative (EC REP or EU AR). For manufacturers outside the EU, appointing one is a legal requirement – and a key partner for compliance and risk management. This article covers the definition, duties, value and selection criteria.
1. Who is the authorised representative?
Under the EU Medical Device Regulation (MDR), a manufacturer not established in the EU may only place devices on the EU market if it has designated a single authorised representative established in the Union. The AR is:
- a party sharing legal responsibility: it performs regulatory tasks on behalf of the manufacturer, such as verifying compliance documents and cooperating on vigilance;
- the regulatory contact point: the official contact for competent authorities in the EU;
- a coordinator in market surveillance: it supports recalls, field safety notices and other urgent measures.
2. What the regulations say
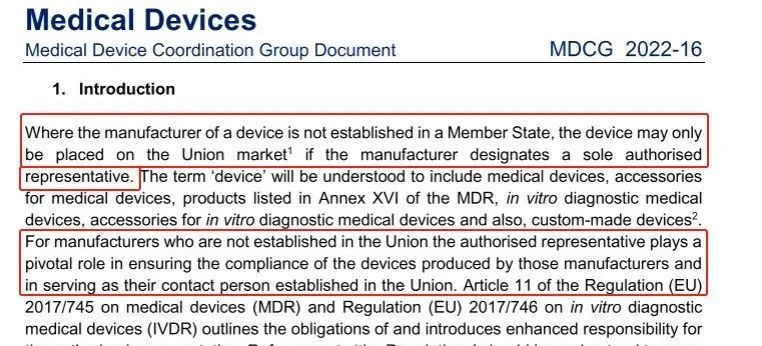
MDCG 2022-16, Section 1: if the manufacturer of a device is not established in a Member State, the device may only be placed on the EU market if the manufacturer designates a sole authorised representative. The AR plays a key role in ensuring the compliance of devices produced by such manufacturers and in serving as their contact person in the EU.
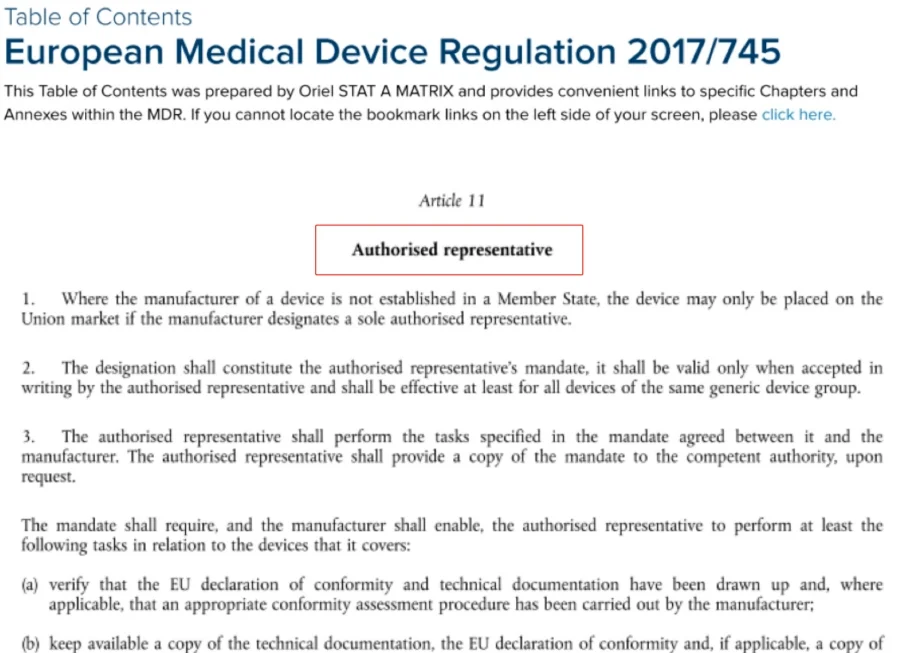
MDR Article 11: for non-EU manufacturers the AR is a precondition for market access. Its tasks include verifying the declaration of conformity and technical documentation, keeping documents available for the authorities, verifying and complying with registration obligations, responding to requests from competent authorities, providing samples or access to the device, cooperating on corrective and preventive action, forwarding complaints and incident reports, and terminating the mandate if the manufacturer acts contrary to its obligations.
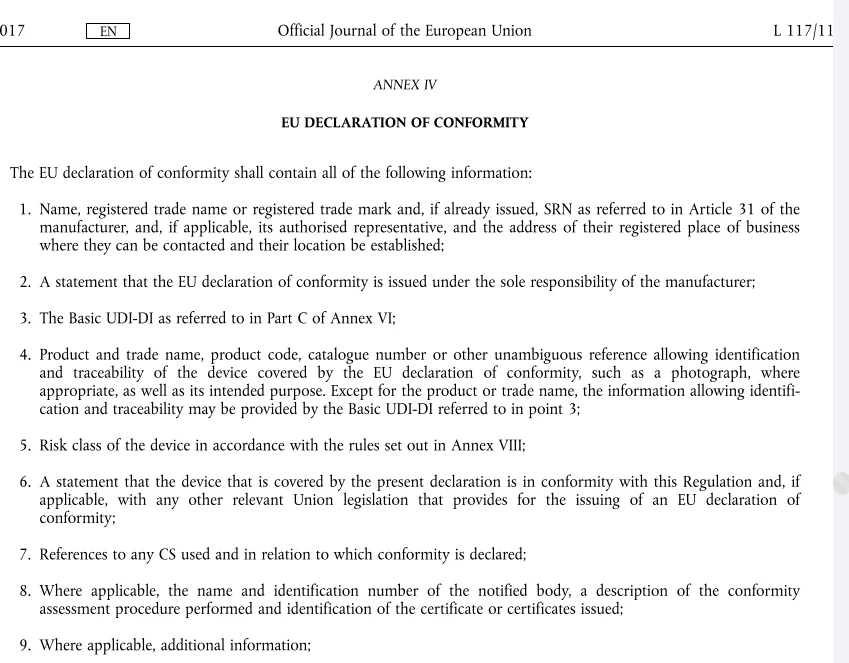
MDR Annex IV, point 1: the EU declaration of conformity must contain the name, registered trade name or registered trade mark and, if already issued, the SRN of the manufacturer and, if applicable, of its authorised representative, together with the address of the registered place of business where they can be contacted.
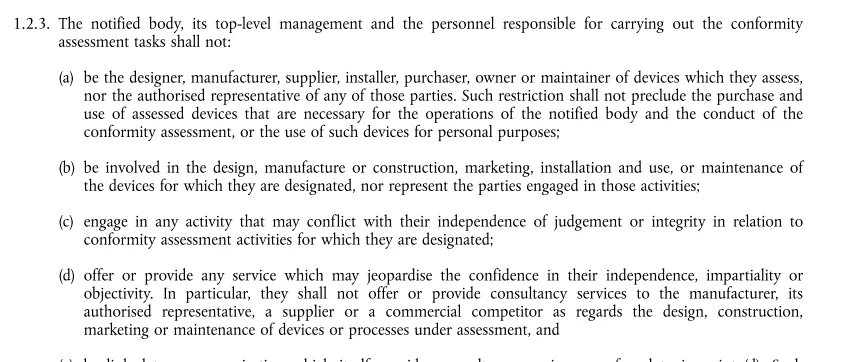
MDR Annex VII (requirements for notified bodies): a notified body may not be the designer, manufacturer, supplier, installer, purchaser, owner or maintainer of the devices it assesses, nor the authorised representative of any of those parties, and it may not provide consultancy services to the manufacturer or its authorised representative concerning the design, construction, marketing or maintenance of the devices assessed.
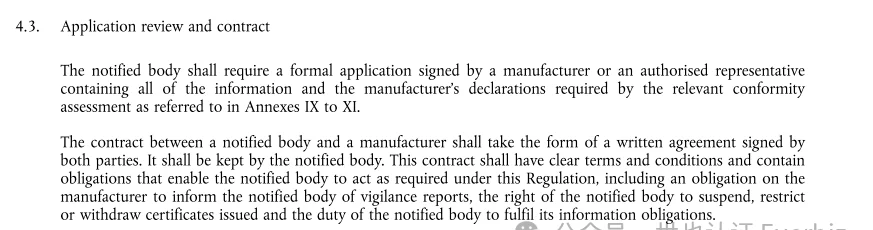
When reviewing an application, the notified body requires a formal application lodged by the manufacturer or its authorised representative.
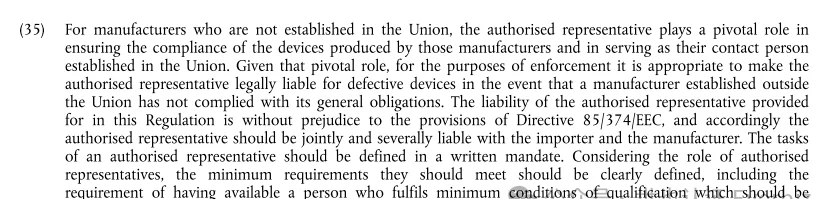
MDR recital 35: for manufacturers outside the Union the authorised representative plays a pivotal role in ensuring compliance and acts as their contact person in the EU. Where the manufacturer has not complied with its obligations, the authorised representative is jointly and severally liable with the importer and the manufacturer for defective devices. Its tasks must be defined in a written mandate, and it must meet minimum requirements, including qualified personnel.

The manufacturer must ensure that the authorised representative permanently has the documentation it needs to perform the tasks in Article 11(3).
3. Why an AR is valuable
- Managing compliance risk: authorities contact the AR, which helps resolve compliance questions quickly and reduces the manufacturer’s direct exposure.
- Faster market access: an experienced AR knows the EU procedures and helps prepare documentation and coordinate with notified bodies.
- Credibility: the qualifications and reputation of the AR influence how authorities and customers perceive the product.
- Long-term compliance: the AR monitors regulatory changes and supports the manufacturer during market surveillance and audits.
4. Five criteria for choosing an AR
- Expertise: medical device background and solid knowledge of the MDR, IVDR and classification, especially for higher-risk devices.
- Experience and set-up: experience with similar products and correct registration as an actor (SRN) in EUDAMED.
- Location and network: a base in a major market such as Germany helps with on-site communication and quick responses.
- Languages and communication: fluent English and the relevant EU languages (e.g. German) to avoid misunderstandings.
- Scope and fees: clearly defined services (e.g. post-market support, crisis management) and transparent pricing without hidden costs.
5. Example: AR support for a wound dressing
A Chinese manufacturer of a chitosan haemostatic dressing needed EU market access. Because the dressing is intended for wounds that breach the dermis, it fell into Class IIb and required robust clinical evidence. The AR provided regulatory support on classification, biocompatibility and technical documentation requirements, handled post-market incident communication with authorities in several Member States, and monitored market feedback to improve labelling in German and Italian.
6. Outlook
- Digital supervision: with EUDAMED becoming mandatory, ARs verify and support registrations in the database.
- Supply-chain transparency: ARs help manufacturers manage UDI traceability.
- Rising compliance effort: more small and medium-sized companies will rely on professional AR services.
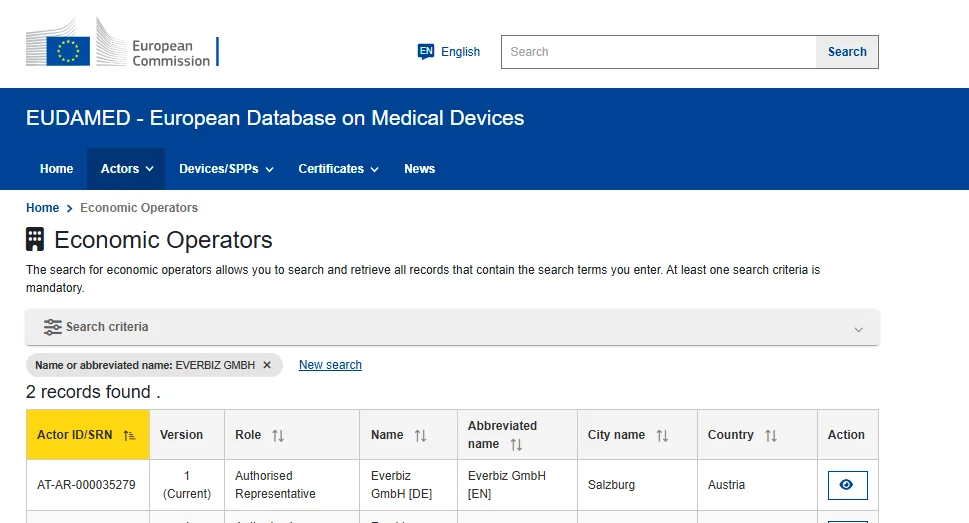
How EVERBIZ can help
EVERBIZ acts as EU authorised representative with registrations in Germany and Austria and can register products under either SRN. We keep the required documents permanently available, liaise with notified bodies and authorities, and offer support in Chinese, English and German. Contact us to discuss a mandate.
This article is for general information only and does not constitute legal advice. The official EU legal texts are authoritative.